• Skip to primary navigation
  • Skip to main content
  • Skip to primary sidebar
  • Skip to footer

MassDevice

The Medical Device Business Journal — Medical Device News & Articles | MassDevice

  • Special Reports
  • Technologies
    • Artificial Intelligence (AI)
    • Cardiovascular
    • Orthopedics
    • Neurological
    • Diabetes
    • Surgical Robotics
  • Business & Finance
    • Wall Street Beat
    • Earnings Reports
    • Funding Roundup
    • Mergers & Acquisitions
    • Initial Public Offering (IPO)
    • Legal News
    • Personnel Moves
    • Medtech 100 Stock Index
  • Regulatory
    • Food & Drug Administration (FDA)
    • Recalls
    • 510(k)
    • Pre-Market Approval (PMA)
    • MDSAP
    • Clinical Trials
  • Resources
    • About MassDevice
    • Leadership in Medtech
    • Manufacturers & Suppliers Search
    • MedTech100 Index
    • Videos
    • Webinars
    • Whitepapers
    • Voices
    • In-Depth Coverage
    • Latest News
  • Attend DeviceTalks
    • Events
      • DeviceTalks Minnesota – May 4
      • DeviceTalks Boston – May 27–28
      • DeviceTalks West
      • DeviceTalks Tuesdays
    • DeviceTalks Podcast Network
      • DeviceTalks Weekly
      • AbbottTalks
      • Boston ScientificTalks
      • DeviceTalks AI
      • IntuitiveTalks
      • MedtechWOMEN Talks
      • MedtronicTalks
      • Neuro Innovation Talks
      • Ortho Innovation Talks
      • Structural Heart Talks
      • StrykerTalks
  • Advertise
  • Subscribe
Home » Artivion earns FDA nod for aortic arch prosthesis device

Artivion earns FDA nod for aortic arch prosthesis device

June 29, 2026 By Sean Whooley

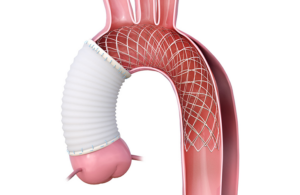
Artivion AMDS hybrid prosthesis
The AMDS hybrid prosthesis device [Image courtesy of Artivion]
Artivion (NYSE:AORT) announced today that it received FDA premarket approval (PMA) for its AMDS Hybrid Prosthesis device.

The approval covers acute Debakey Type 1 aortic dissections with either clinical or radiographic malperfusion. Atlanta-based Artivion estimates that this population comprises approximately 60% of all Debakey Type 1 aortic dissections. The condition affects approximately 6,000 patients in the U.S. each year, the company says. When left untreated, mortality comes in at approximately 1% per hour and up to 50% within the first 48 hours.

Previously, the device held FDA humanitarian device exemption (HDE). Now, hospitals no longer require institutional review board approval (IRB) to implant AMDS. This reduces administrative burden and enables more immediate treatment.

Artivion labels the device as the the world’s first aortic arch remodeling device for use in the treatment of acute DeBakey Type I aortic dissections. The company says it helps to significantly reduce the incidence of distal anastomotic new entry (DANE) tears. It complements hemiarch replacement without adding technical complexity when deployed in the aortic arch during a standard ascending aorta replacement. The device preserves the native arch, enabling minimally invasive reinterventions if needed.

FDA approval came on the heels of data from the PERSEVERE U.S. IDE trial. At 30 days, the study demonstrated a 72% reduction in all-cause mortality and a 54% reduction in primary major adverse events.

Pat Mackin, chair, president and CEO of Artivion, said:

“We are thrilled to receive AMDS PMA approval, as it not only validates the enduring benefits shown in the PERSEVERE clinical data but also removes a barrier to broader adoption by eliminating the IRB requirement that came with the HDE. Having already built commercial and clinical infrastructure during our HDE launch, and with strong reordering momentum among accounts already using AMDS, we expect this PMA to help accelerate adoption of this life saving technology.

“This approval, following our recent acquisition of the PMA-approved NEXUS system and continued clinical trial enrollment progress for ARCEVO LSA, marks another step forward in our vision of Artivion as the only company globally with a complete portfolio of market leading aortic arch solutions. We are deeply grateful to every PERSEVERE investigator and patient who helped make this possible.”

Filed Under: Endoscopic, Food & Drug Administration (FDA), Implants, Pre-Market Approval (PMA), Regulatory/Compliance, Stent Grafts, Stents, Vascular Tagged With: Artivion, FDA

More recent news

  • Former Shockwave Medical CEO joins Solenic Medical’s board
  • Anaconda Biomed brings in $56 million
  • Precision Neuroscience raises $250M in Series D for its BCI
  • Fresenius discontinuing Chinese sales of 4008A hemodialysis system
  • Perfuze’s super-bore stroke catheter tops conventional, cyclic aspiration in bench study

About Sean Whooley

Sean Whooley is a senior editor who mainly produces work for MassDevice, Medical Design & Outsourcing and Drug Delivery Business News. He received a bachelor's degree in multiplatform journalism from the University of Maryland, College Park. You can connect with him on LinkedIn or email him at [email protected].

Primary Sidebar

Cart

“md
EXPAND YOUR KNOWLEDGE AND STAY CONNECTED
Get the latest med device regulatory, business and technology news.

DeviceTalks Weekly

See More >
MDO ad
MDO ad

Footer

MASSDEVICE MEDICAL NETWORK

DeviceTalks
Drug Delivery Business News
Medical Design & Outsourcing
Medical Tubing + Extrusion
Drug Discovery & Development
Pharmaceutical Processing World
MedTech 100 Index
R&D World
Medical Design Sourcing

DeviceTalks Webinars, Podcasts, & Discussions

Attend our Monthly Webinars
Listen to our Weekly Podcasts
Join our DeviceTalks Tuesdays Discussion

MASSDEVICE

Subscribe to MassDevice E-Newsletter
Advertise with us
About
Contact us

Copyright © 2026 · Arrowfly LLC and its licensors. All rights reserved.
The material on this site may not be reproduced, distributed, transmitted, cached or otherwise used, except with the prior written permission of Arrowfly.

Privacy Policy