Amsterdam-based Philips’ testing showed that, among other things, exposure to certain emissions from the recalled devices is “unlikely to result in appreciable harm to health in patients.”
The update marks the latest development in the long recall saga surrounding the company’s Philips Respironics subsidiary.
Last year, Philips stopped taking orders of sleep therapy systems as it handled a Class I recall of certain devices. Respironics recalled millions of ventilators, bi-level positive airway pressure (BiPAP) and continuous positive airway pressure (CPAP) machines.
It attributed the recall to potential health risks caused by polyester-based polyurethane (PE-PUR) sound abatement foam breaking down. Foam particles could enter a device’s air pathway. This could cause a range of potential health problems and toxic, carcinogenic effects for the user.
The most recent update, issued by the FDA last month, said the agency received more than 90,000 reports of problems — including 260 mentioning deaths.
Philips says testing showed unlikely harm from devices
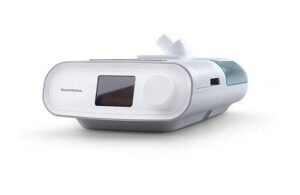
First-generation DreamStation devices assessed by Philips comprise approximately 68% of the registered devices globally. Those devices were not exposed to ozone cleaning in line with instructions for use.
Philips conducted its testing with five independent certified testing laboratories. The company, along with third-party qualified experts and an external panel, reviewed and assessed results.
New results feature testing on exposure to particulate matter (PM) emissions from degraded foam in DreamStation devices. That includes potential respirable and non-respirable particulates. Results demonstrated these emissions are unlikely to result in appreciable harm to health. PM emissions from used devices with degradation proved not statistically different than those form devices without degradation. Philips said this indicates that degradation did not contribute to appreciable elevated levels of respirable particles in tested devices.
Additionally, Philips’ testing evaluated exposure to volatile organic compound emissions (VOCs). Results showed that this “is not anticipated to result in long-term health consequences for patients.”
Finally, testing found the prevalence of visible foam degradation in inspected, returned devices to be low. In total, 164 out of 36,341 (0.5%) returned devices from the U.S. and Canada showed significant visible degradation. One out of 2,469 (0.04%) devices returned from Europe showed the visible degradation. Philips said this falls in line with results it presented in June.
Ozone cleaning results
Philips also reported data demonstrating that ozone cleaning exacerbates foam degradation. Returned devices from the U.S. and Canada demonstrated a 14-times higher likelihood of visible foam degradation compared to those with no user-reported ozone exposure.
Third-party data concluded that exposure to particulates from degraded foam with self-reported ozone use in first-generation DreamStation devices is “unlikely to result in appreciable harm to health in patients.”
Philips said the VOC toxicological risk of ozone-induced foam degradation remains under evaluation.
Philips leadership speaks about the update
Roy Jakobs took over as Philips CEO in the midst of the recall in October. At the time of his appointment, he apologized for the company’s ongoing issues.
In the company’s update today, he said the company’s priority remains the health and well-being of patients. He again apologized, saying the company is sorry for the amount of time its assessments of the situation have taken. He said testing involved long throughput times and required thoroughness.
“We deeply regret the concern and uncertainty felt by patients, their physicians, and customers, and continue to work hard to address their needs,” Jakobs said. “By year-end, we have ramped up our production such that we have produced almost 90% of all devices that are needed for the remediation program. The relevant competent authorities globally, including the FDA, are still reviewing the extensive data and assessments that we have provided. We share the same objective to ensure patient safety and quality in the delivery of healthcare. We are therefore committed to working closely with these agencies, as we continue to complete the test and research program, and the remediation of the affected devices.”
The history behind the recall
Following its June 2021 voluntary recall notice, Philips dropped out of the sleep therapy market. It has yet to return.
In March 2022, the FDA told Philips that its notification efforts on the recall of its ventilators had been inadequate to that point. A month later, the company revealed a Department of Justice subpoena for information related to the recall. The subpoena sought “information related to events leading to the Respironics recall.” Philips said its subsidiaries are cooperating with investigators.
On May 2, the FDA’s Center for Devices and Radiological Health (CDRH) proposed an order for Philips to submit a plan for the repair, replacement or refund of the purchase price of the recalled devices. The order would cover devices manufactured after November 2015.
Also in May, the FDA confirmed that it received more than 21,000 medical device reports (MDRs), including 124 reports of death, between April 2021 and April 30, 2022. It later reported 48,000 MDRs and 44 reports of death between May 1 and July 31 of this year.
In August, an FDA update brought the death total to 168, with 69,000 MDRs filed related to the recall. The latest number reported is the aforementioned 260 deaths on more than 90,000 MDRs. (Note: The FDA’s Medical Device Reporting (MDR) system remains a passive system with limitations.)
Philips Respironics said it continues to remediate devices affected by the recall. It expects to complete 90% of the production this year for the delivery of replacement devices to patients.